Last Updated: August 7, 2026
Peptide quality standards for clinical trials represent the comprehensive set of specifications, analytical requirements, and regulatory compliance measures that ensure therapeutic peptides meet safety, purity, and efficacy benchmarks before human testing. A contaminated or degraded peptide batch can invalidate months of research, compromise patient safety, or trigger regulatory rejection at submission.
At Rejuvenation Wellness Solutions LLC, we understand that researchers conducting clinical trials demand peptides that meet these exacting standards. Our partnership with WHO/GMP and ISO-certified manufacturers ensures that every batch undergoes rigorous independent lab testing with a Certificate of Analysis, confirming 99%+ purity before reaching your laboratory.
The regulatory landscape combines FDA’s current Good Manufacturing Practice (cGMP) requirements with pharmacopeial standards from the United States Pharmacopeia (USP) and European Pharmacopoeia (EP). This guide walks you through each component of peptide quality standards, from FDA cGMP requirements to supply chain integrity measures that protect your trial data from contamination at every stage.
The FDA’s Current Good Manufacturing Practice regulations apply directly to synthetic peptides intended for clinical trials. These requirements mandate that manufacturers establish and maintain quality control strategies ensuring consistent batch-to-batch purity, identity, and potency throughout the manufacturing process.
cGMP compliance means implementing written procedures for every manufacturing step: raw material qualification, synthesis conditions, purification methods, and final product testing. Manufacturers must document standard operating procedures (SOPs) for equipment maintenance, environmental monitoring, and personnel training. Deviations must trigger investigation and corrective action.
For peptides specifically, cGMP requires manufacturers to establish acceptance criteria based on the drug’s intended use. A peptide destined for a Phase I trial may have different purity thresholds than one advancing to Phase III. The FDA expects manufacturers to justify these criteria through preclinical data, stability studies, and analytical method validation.
cGMP mandates that manufacturers retain comprehensive batch records including raw material certificates of analysis, in-process testing results, final product specifications, and environmental monitoring data. If your trial faces an FDA audit or adverse event investigation, these records become central to demonstrating that your peptide met established quality standards at manufacture.
Peptide impurity profile analysis identifies and quantifies all detectable impurities in a batch, from unreacted starting materials and synthesis intermediates to degradation products and process-related contaminants. Acceptance criteria define the maximum allowable levels for each impurity class.
The impurity profile typically includes peptide-related impurities (truncated sequences, misfolded conformers, oxidized variants), process-related impurities (residual solvents, reagent residues, salts), and inorganic impurities (heavy metals, endotoxins).
Acceptance criteria for therapeutic peptides generally follow a tiered approach. The primary peptide sequence must comprise at least 95-99% of total peptide content, depending on therapeutic class and development phase. Individual impurities above a certain threshold (typically 0.05-0.10%) must be identified and quantified. Impurities below the quantitation limit but above the detection limit are reported as "detected but not quantified."
The challenge lies in setting realistic acceptance criteria. Set them too tight, and you’ll reject batches that are actually safe and effective. Set them too loose, and you risk undetected degradation that could compromise your trial. The FDA expects manufacturers to establish criteria based on analytical data from stability studies, toxicology assessments, and clinical experience with similar peptides.
Analytical methods form the backbone of peptide quality assessment. They must be validated, reproducible, and capable of detecting impurities at clinically relevant levels. The primary methods include high-performance liquid chromatography (HPLC), mass spectrometry (MS), and amino acid analysis.
Reverse-phase HPLC separates peptide components based on their chemical properties, allowing quantification of the primary peptide and resolution of impurities. The method must be validated to demonstrate specificity, linearity, accuracy, and precision across the expected concentration range.
Mass spectrometry determines the exact molecular weight of the peptide and identifies structural variants. Electrospray ionization (ESI) mass spectrometry is particularly useful because it can analyze peptides in solution without requiring derivatization. Time-of-flight (TOF) analyzers provide high-resolution mass measurements that can distinguish between peptides differing by a single amino acid.
Amino acid analysis quantifies the elemental composition after acid hydrolysis, confirming that the peptide contains the expected amino acids in the correct ratios and validating the primary sequence indirectly.
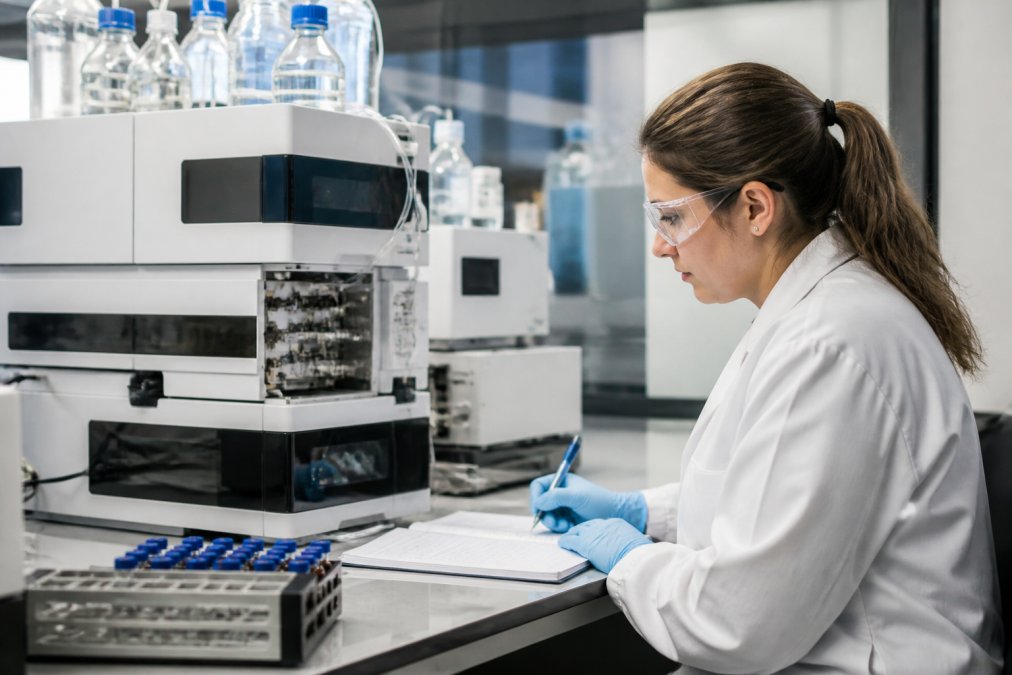
Beyond these primary methods, peptide characterization may include peptide sequence verification through Edman degradation, secondary structure assessment via circular dichroism spectroscopy, and aggregation state assessment through size-exclusion chromatography. All analytical methods used to support clinical trial submissions must be validated according to ICH Q2(R2) guidelines.
Stability testing establishes how peptide quality changes over time under defined storage conditions. These data are essential for setting expiration dates and establishing storage requirements that maintain quality throughout your clinical trial.
Stability protocols typically follow ICH Q1A(R2) guidelines, which define three standard storage conditions: long-term storage at 25°C ± 2°C / 60% RH ± 5% RH (12 months minimum), intermediate storage at 30°C ± 2°C / 75% RH ± 5% RH (6 months minimum), and accelerated storage at 40°C ± 2°C / 75% RH ± 5% RH (3-6 months minimum).
Testing intervals are front-loaded: samples are analyzed at baseline, then at 1, 2, 3, and 6 months for accelerated conditions. For long-term storage, testing typically occurs at 3, 6, 9, and 12 months. Each timepoint measures assay (peptide content), impurity profile, moisture content, and pH (if applicable).
The data reveal degradation pathways. Peptides may oxidize at elevated temperatures, aggregate under certain humidity conditions, or hydrolyze if exposed to moisture. Understanding these pathways allows you to establish storage conditions that slow degradation and extend shelf life.
For clinical trials specifically, stability data must support the timeframe of your study. If your trial spans 18 months from manufacture to final patient assessment, you need stability data demonstrating that the peptide remains within specification for at least that duration.
Supply chain integrity ensures that your peptide maintains its quality from manufacture through final use in your trial. Contamination, degradation, or substitution at any point can invalidate your research and compromise participant safety.
Vendor qualification begins with assessing the manufacturer’s quality systems. Request their quality manual and evidence of regulatory inspections. A manufacturer with a clean inspection history or documented resolution of past findings demonstrates commitment to quality.
Evaluate your shipping and storage partner carefully. Temperature excursions during transit can degrade peptides, particularly those sensitive to heat or moisture. Require validated shipping containers with temperature monitoring and documentation of shipping conditions.
Upon receipt, implement receiving inspection procedures. Verify that the shipment arrived at the correct temperature, that vials show no visible signs of damage, and that the Certificate of Analysis matches the batch number on the vials.
Rejuvenation Wellness Solutions LLC prioritizes supply chain integrity through partnerships with WHO/GMP and ISO-certified manufacturers. Every batch undergoes independent lab testing with a Certificate of Analysis, and we offer same-day order fulfillment with UPS 2-day air shipping to minimize time in transit.
For long-term trials, implement periodic re-testing of stored peptide supplies. Testing samples from your inventory at defined intervals (typically annually) confirms that your storage conditions are adequate and alerts you to unexpected degradation.
The United States Pharmacopeia (USP), European Pharmacopoeia (EP), and Japanese Pharmacopoeia (JP) each publish standards for peptide quality. While these standards converge on core principles, differences in methodology and acceptance criteria can create compliance challenges for trials spanning multiple regulatory jurisdictions.
USP standards for peptides emphasize identity confirmation through mass spectrometry or amino acid analysis, purity assessment via HPLC, and characterization of the primary sequence. Acceptance criteria in USP monographs typically require 95% minimum assay for the primary peptide.
The European Pharmacopoeia applies similar principles but sometimes specifies different analytical methods or acceptance ranges. These variations, while often minor, can create compliance complications if your trial operates under both US and European regulatory oversight.
The International Council for Harmonisation (ICH) has published Q3B(R2) guidance on impurities in new drug substances, providing a framework for harmonizing these standards. For multinational trials, the practical approach is to establish specifications that satisfy the most stringent requirement across all applicable pharmacopeias.
Regulatory submissions for peptide-based clinical trials require comprehensive documentation of quality systems, analytical methods, and manufacturing processes. Your submission must include drug substance characterization, analytical method validation, stability data, manufacturing process description, and quality control strategy.
The validation section is particularly critical. Regulators scrutinize analytical method validation to ensure that your testing actually measures what you claim it measures and that results are reproducible across different analysts and instruments.
For peptides, the FDA also expects discussion of peptide-related impurities, truncated sequences, misfolded variants, and oxidized forms. You must demonstrate that you’ve identified these impurities, established methods to detect them, and set acceptance criteria that protect clinical trial participants.
Documentation must be contemporaneous. All records, analytical reports, and batch records must be dated and signed at the time of testing. Retrospectively compiled documentation raises red flags with regulators.
Rejuvenation Wellness Solutions LLC supports researchers navigating these requirements by providing comprehensive Certificates of Analysis that document all testing performed on your peptide batch. Our independent lab testing includes the analytical data needed for regulatory submissions, reducing the burden on your team to conduct duplicate testing.
| Documentation Element | Requirement | Purpose |
|---|---|---|
| Analytical Method Validation | ICH Q2(R2) compliant validation reports | Demonstrates method suitability for regulatory use |
| Batch Records | Complete manufacturing records with dates and signatures | Establishes manufacturing history and quality control |
| Stability Data | Long-term, intermediate, and accelerated studies | Supports shelf life and storage conditions |
| Impurity Characterization | Identified and quantified impurities with structure elucidation | Confirms safety profile of the peptide |
| Certificate of Analysis | Summary of all testing with results and acceptance criteria | Provides rapid verification of batch quality |
Peptide quality standards for clinical trials represent the convergence of rigorous science, regulatory compliance, and practical laboratory operations. The manufacturers and suppliers you choose directly impact your ability to meet these standards. Rejuvenation Wellness Solutions LLC partners with WHO/GMP and ISO-certified facilities to deliver peptides that meet these exacting requirements, with independent lab testing and same-day fulfillment to support your research timeline. Shop with confidence knowing your peptides have been tested and verified to meet clinical-grade quality standards.
The FDA requires that peptide reference standards meet strict purity thresholds, typically 95% or higher depending on the therapeutic peptide's intended use. Manufacturers must follow FDA cGMP (current Good Manufacturing Practice) guidelines and provide comprehensive analytical data demonstrating purity through methods like HPLC and mass spectrometry. Every batch destined for clinical trials requires a Certificate of Analysis documenting the purity profile and absence of harmful impurities. The specific acceptance criteria depend on the peptide's pharmacological role and must be established during drug development and approved in regulatory submissions.
Impurity profiling identifies and quantifies all chemical and biological contaminants in a peptide batch, which directly affects safety and efficacy in human studies. Even small amounts of impurities can trigger immune responses, reduce bioavailability, or introduce unexpected toxicity. Regulatory agencies require detailed impurity characterization to establish acceptance criteria and understand potential risks to trial participants. Synthetic peptides often contain residual solvents, process-related impurities, and degradation products that must be controlled and documented. Without rigorous impurity analysis, clinical trials cannot proceed because the true composition and safety profile of the therapeutic peptide remains unknown.
Regulatory submissions require comprehensive manufacturing documentation including batch records, analytical test results, stability data, process validation reports, and quality control strategies. Each batch must include a Certificate of Analysis showing purity, potency, identity verification through amino acid analysis or peptide sequence verification, and testing for residual solvents and endotoxins. Manufacturers must document their GMP compliance, equipment qualification, and vendor qualification records for raw materials. For clinical trials, this documentation supports the Investigational New Drug (IND) application and demonstrates that every batch meets the same specifications and quality standards required for human use.
Validation involves establishing and documenting that your analytical methods reliably measure peptide purity, identity, and potency. You must perform method validation studies proving accuracy, precision, and specificity for your reference standards and acceptance criteria. Stability testing protocols assess how the peptide degrades under storage conditions to set realistic shelf-life claims. Batch consistency testing confirms that multiple production runs meet the same specifications. All validation data, analytical methods, and acceptance criteria must be submitted to regulatory bodies and approved before the first clinical dose is administered. This ensures that every batch used in the trial meets the validated quality profile.
This article was written using GrandRanker